


This is the mapper pipeline from the Sequana projet
- Overview:
This is a simple pipeline to map several FastQ files onto a reference using different mappers/aligners
- Input:
A set of FastQ files (illumina, pacbio, etc).
- Output:
A set of BAM files (and/or bigwig) and HTML report
- Status:
Production
- Documentation:
This README file, and https://sequana.readthedocs.io
- Citation:
Cokelaer et al, (2017), ‘Sequana’: a Set of Snakemake NGS pipelines, Journal of Open Source Software, 2(16), 352, JOSS DOI https://doi:10.21105/joss.00352
Installation
If you already have all requirements, you can install the packages using pip:
pip install sequana_mapper --upgrade
You will need third-party software such as bwa and samtools. Please see below for details.
Usage
Scan FastQ files in a directory and set up the pipeline (replace DATAPATH and genome.fa with your inputs):
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --aligner-choice bwa
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --aligner-choice bwa --do-coverage
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --aligner-choice bwa --create-bigwig
For long-read data, use the dedicated presets:
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --pacbio     # sets minimap2 -x map-pb
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --nanopore   # sets minimap2 -x map-ont
For capture-seq projects (feature counting):
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --capture-annotation-file targets.saf
This creates a mapper/ directory with the pipeline and configuration file. Execute the pipeline locally:
cd mapper
bash mapper.sh
See .sequana/profile/config.yaml to tune Snakemake behaviour (cores, cluster settings, etc.).
Usage with apptainer
With apptainer, initiate the working directory as follows:
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --use-apptainer
Images are downloaded in the working directory but you can store them in a shared location:
sequana_mapper --input-directory DATAPATH --reference-file genome.fa --use-apptainer --apptainer-prefix ~/.sequana/apptainers
and then:
cd mapper
bash mapper.sh
Requirements
This pipeline requires the following executables (install via bioconda/conda):
bwa — short-read aligner (default)
minimap2 — long-read aligner (PacBio / Nanopore)
bowtie2 — alternative short-read aligner
samtools / sambamba — BAM processing
bamtools — BAM statistics
fastqc — raw-read quality control (optional, --do-fastqc)
deeptools — bigwig generation (bamCoverage)
bedtools — genome arithmetic
subread — feature counting (featureCounts, capture-seq only)
mosdepth — fast coverage depth
seqkit — FASTQ statistics
multiqc — aggregated HTML report
sequana_coverage — coverage analysis (prokaryotes)
Install all dependencies at once:
mamba env create -f environment.yml
Details
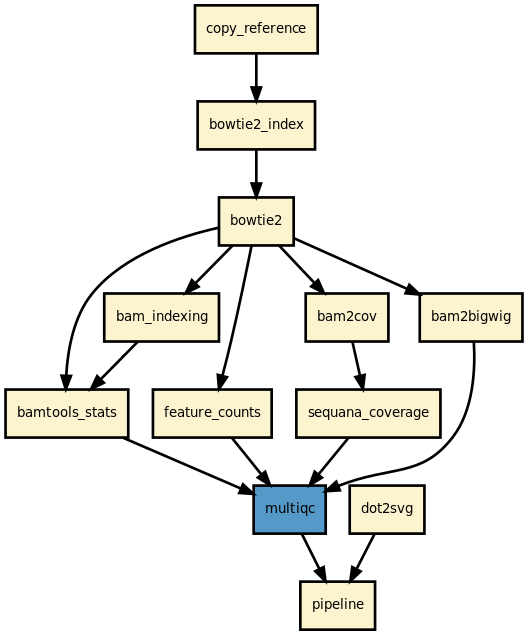
This pipeline maps FastQ files (paired or single-end) in parallel onto a reference genome and produces filtered BAM files, a MultiQC HTML report, and optionally coverage tracks and feature counts.
Aligner choice (--aligner-choice):
bwa (default) — BWA-MEM; index algorithm is auto-selected (is or bwtsw) based on reference size
bwa_split — experimental; splits large FastQs into 1 M-read chunks for parallel BWA jobs, then merges
minimap2 — long-read aligner; use --pacbio (sets -x map-pb) or --nanopore (sets -x map-ont)
bowtie2 — standard short-read aligner
BAM filtering: unmapped reads are removed to minimise file size. Statistics reported by MultiQC (in {sample}/bamtools_stats/) still include both mapped and unmapped read counts.
Optional outputs:
--do-coverage — runs sequana_coverage for depth-of-coverage analysis (prokaryotes)
--do-fastqc — runs fastqc on the raw input reads (included in the MultiQC report)
--create-bigwig — generates bigwig files via bamCoverage (deeptools)
--capture-annotation-file — enables featureCounts for capture-seq efficiency metrics
Rules and configuration details
Here is the latest documented configuration file to be used with the pipeline. Each rule used in the pipeline may have a section in the configuration file.
Changelog
Version |
Description |
|---|---|
1.5.0 |
|
1.4.1 |
|
1.4.0 |
|
1.3.1 |
|
1.3.0 |
|
1.2.1 |
|
1.2.0 |
|
1.1.0 |
|
1.0.0 |
|
0.12.0 |
|
0.11.1 |
|
0.11.0 |
|
0.10.1 |
|
0.10.0 |
|
0.9.0 |
|
0.8.13 |
|
0.8.12 |
|
0.8.11 |
|
0.8.10 |
|
0.8.9 |
|
0.8.8 |
|
0.8.7 |
|
0.8.6 |
|
0.8.5 |
|
0.8.4 |
|
0.8.3 |
|
0.8.2 |
|
0.8.1 |
|
0.8.0 |
First release. |
Contribute & Code of Conduct
To contribute to this project, please take a look at the Contributing Guidelines first. Please note that this project is released with a Code of Conduct. By contributing to this project, you agree to abide by its terms.
1 maintainer

Metadata
1 maintainer
Release files for sequana-mapper 1.5.0
For a detailed explanation of source distributions (sdists) and built distributions (wheels), please see the package formats documentation.
Source distribution (sdist)
| File | Size | Uploaded | |
|---|---|---|---|
| sequana_mapper-1.5.0.tar.gz | 74.2 kB | Details |
Built distribution (wheel)
| File | Interpreter | ABI | Platform | Reset |
|---|---|---|---|---|
| sequana_mapper-1.5.0-py3-none-any.whl | Python 3 | none | any | Details |
Total release size: 146.8 kB
Release files / sequana_mapper-1.5.0.tar.gz
| Download URL | sequana_mapper-1.5.0.tar.gz |
|---|---|
| Size | 74.2 kB |
| Tags | Source |
|
SHA-256 checksum How to use checksums |
68265dc658d35ff4e352b32d3dcec9d6e56d3694dc4c435d7cb9869dc613f133
|
|
BLAKE2b-256 checksum How to use checksums |
8ee7622f772a882e5af9dd1a80ffde77e5c8188cc0e2cd252f94d91b8b942de2
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
No |
| Uploaded via |
twine/6.1.0 CPython/3.13.12
|
Release files / sequana_mapper-1.5.0-py3-none-any.whl
| Download URL | sequana_mapper-1.5.0-py3-none-any.whl |
|---|---|
| Size | 72.6 kB |
| Tags | Python 3 |
|
SHA-256 checksum How to use checksums |
1055442b43a16fd7db3daccfa5f3f18d08bceefaec35f52cccda8f7fa5f6a928
|
|
BLAKE2b-256 checksum How to use checksums |
f681e3165994e16acea376b20e56dbdd24507bc2e1536b0b8fe5f39f3be57e58
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
No |
| Uploaded via |
twine/6.1.0 CPython/3.13.12
|














