
MethylSeg
MethylSeg is a Python toolkit for identifying methylation domains from whole-genome bisulfite sequencing (WGBS) and microarray methylation data. It supports data preparation, methylation-state model training, genome segmentation, region cleaning, and visualization.
This introduces the standard workflow. For the complete API reference and example notebooks, see the MethylSeg documentation.
Hardware requirements
Memory usage depends on the input data type and dataset size. The following values were observed during testing:
| Data type | Typical memory usage | Recommended system memory |
|---|---|---|
| WGBS | ~12 GB average; up to ~15 GB peak | ≥16 GB |
| HM450K | <1 GB | ≥8 GB |
Installation
Install from PyPI
python -m pip install methylseg
or install the current version from GitHub:
python -m pip install "git+https://github.com/clementlab/MethylSeg.git"
Reference files
Sample data
Three reference samples are available for use in tutorials and testing.
methylseg download_data_files
The two colon cancer samples were originally generated by Hansen et al. (2011) and were obtained from the data distributed with MethyLasso (Balaramane et al., 2024).
The liver cancer sample, TCGA-BD-A3EP-01A, was obtained from the TCGA-LIHC project through the NCI Genomic Data Commons.
| Sample name | WGBS filename | HM450K filename |
|---|---|---|
WGBS_colon-primary-tumor_1 |
WGBS_colon-primary-tumor_1_wgbs.tsv |
WGBS_colon-primary-tumor_1_450k.beta |
WGBS_colon-primary-tumor_2 |
WGBS_colon-primary-tumor_2_wgbs.tsv |
WGBS_colon-primary-tumor_2_450k.beta |
TCGA-BD-A3EP-01A |
N/A | TCGA-BD-A3EP-01A_450k.tsv |
References
- Hansen, K. D., et al. Increased methylation variation in epigenetic domains across cancer types. Nature Genetics 43, 768–775 (2011).
- Balaramane, D., Spill, Y. G., Weber, M. & Bardet, A. F. MethyLasso: a segmentation approach to analyze DNA methylation patterns and identify differentially methylated regions from whole-genome datasets. Nucleic Acids Research 52, e98 (2024).
Pretrained models
Two pretrained models are provided for tutorials, testing, and exploratory analysis:
| Model filename | Training data |
|---|---|
wgbs_colon_model |
WGBS_colon-primary-tumor_1_wgbs.tsv |
tcga_hm450k_model |
TCGA-BD-A3EP-01A_450k.tsv |
Load a pretrained model with get_pretrained_model:
# WGBS model
wgbs_saved = MethylSegPathway.get_pretrained_model(
    out_dir=OUTPUT_DIR,
    resolution="wgbs",
)

# HM450K model
hm450k_saved = MethylSegPathway.get_pretrained_model(
    out_dir=OUTPUT_DIR,
    resolution="450k",
)
Quickstart
The default workflow trains a model on your input, segments the sample, cleans the calls, and draws a cleaned methylation state overlay.
Use resolution="wgbs" for WGBS count tables or resolution="450k" for HM450K
beta-value tables. Other microarray platforms may require manually configured
parameters until array-specific defaults are added.
from pathlib import Path

from methylseg import MethylSegPathway, MethylationStates
from methylseg.helper_classes import DATA_DIR

reference_dir = DATA_DIR / "reference_files"

# WGBS: replace these with your own sample name and count table.
# sample_name = "WGBS_colon-primary-tumor_1"
# sample_file = reference_dir / "WGBS_colon-primary-tumor_1_wgbs.tsv.gz"
# resolution = "wgbs"

# TCGA/HM450K alternative:
sample_name = "TCGA-BD-A3EP-01A"
sample_file = reference_dir / "TCGA-BD-A3EP-01A_450k.tsv.gz"
resolution = "450k"

sample_info, removed_df = MethylSegPathway.prepare_sample_info(
    sample_name=sample_name,
    sample_file=sample_file,
    resolution=resolution,
    remove_low_coverage_like_cpgs=True,
)

pathway = MethylSegPathway(
    train_sample_info=sample_info,
    out_dir=Path("out/quickstart") / sample_name,
)

pathway.run_pathway()

fig = pathway.plot_labels(
    sample_info=sample_info,
    sample_info_removed=removed_df,
    chrom="chr1",
    region_start=2_200_000,
    region_end=3_700_000
)
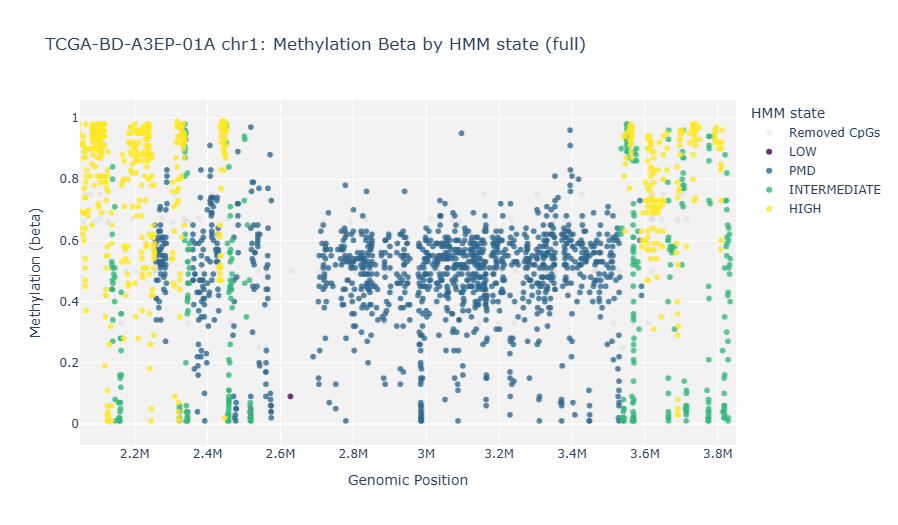
run_pathway(sample_info=sample_info, chroms=["chr1"]) performs fitting,
segmentation, cleaning, and summary-file writing in one call.
Input formats
MethylSeg accepts tab-delimited .tsv and .tsv.gz files containing DNA methylation measurements. Genomic coordinates must use the 0-based, half-open coordinate system, and chromosome names must be consistent throughout each file.
Two BED-like input formats are supported.
Beta-value format
This format can be used for either WGBS or microarray data. It contains four required columns:
- chromosome;
- CpG start position;
- CpG end position; and
- methylation beta value.
For microarray data, an optional fifth column containing the probe identifier may be included. Probe identifiers are retained as metadata.
| CpG_chrm | CpG_beg | CpG_end | beta | probe |
|---|---|---|---|---|
| chr1 | 15864 | 15866 | 0.0 | cg13869341 |
| chr1 | 29406 | 29408 | 0.0 | cg12045430 |
| chr1 | 29424 | 29426 | 0.0 | cg20826792 |
WGBS count format
This format contains five required columns:
- chromosome;
- CpG start position;
- CpG end position;
- methylated read count; and
- total read coverage.
MethylSeg calculates the beta value internally as: $ \mathrm{beta} = \frac{\mathrm{methylated\ reads}}{\mathrm{total\ coverage}}. $
| CpG_chrm | CpG_beg | CpG_end | meth | coverage |
|---|---|---|---|---|
| chr1 | 10468 | 10470 | 14 | 15 |
| chr1 | 10470 | 10472 | 10 | 10 |
| chr1 | 10483 | 10485 | 23 | 28 |
Outputs
The final output of MethylSeg consists primarily of BED files containing the
identified methylation domains. Separate files are generated for each
methylation state: HIGH, INTERMEDIATE, LOW, and PMD.
For each chromosome and state, MethylSeg writes:
| Output | Filename pattern | Description |
|---|---|---|
| Raw regions | segments_{chrom}_{sample_id}_{resolution}_{state}.bed |
Regions produced directly by genome segmentation |
| Cleaned regions | clean_regions/segments_cleaned_{chrom}_{sample_id}_{resolution}_{state}.bed |
Regions retained after merging and filtering |
| Cleaned metadata | clean_regions/metadata_cleaned_{chrom}_{sample_id}_{resolution}_{state}.tsv |
Additional information about the cleaned regions |
After all requested chromosomes have been processed, MethylSeg also creates
genome-wide files for each state under summary_files/:
| Output | Filename pattern |
|---|---|
| Genome-wide raw regions | segments_raw_{state}.bed |
| Genome-wide cleaned regions | segments_cleaned_{state}.bed |
| Genome-wide cleaned metadata | metadata_cleaned_{state}.tsv |
A simplified output directory has the following structure:
methylseg_output/
├── segments_chr1_sample.wgbs_PMD.bed
├── segments_chr1_sample.wgbs_LOW.bed
├── clean_regions/
│   ├── segments_cleaned_chr1_sample.wgbs_PMD.bed
│   ├── segments_cleaned_chr1_sample.wgbs_LOW.bed
│   ├── metadata_cleaned_chr1_sample.wgbs_PMD.tsv
│   └── metadata_cleaned_chr1_sample.wgbs_LOW.tsv
└── summary_files/
    ├── segments_raw_PMD.bed
    ├── segments_raw_LOW.bed
    ├── segments_cleaned_PMD.bed
    ├── segments_cleaned_LOW.bed
    ├── metadata_cleaned_PMD.tsv
    └── metadata_cleaned_LOW.tsv
Each BED file contains 0-based, half-open genomic coordinates and the assigned methylation state. For example, segments_cleaned_PMD.bed may contain:
| Chromosome | Start | End | State |
|---|---|---|---|
chr1 |
1,261,344 | 1,323,691 | PMD |
chr1 |
2,789,167 | 2,999,312 | PMD |
chr1 |
3,885,138 | 3,916,529 | PMD |
chr1 |
6,167,866 | 6,186,177 | PMD |
The BED files themselves are tab-delimited and do not contain a header.
Citation
If you use MethylSeg, please cite the software using the CITATION.cff metadata. On GitHub, select Cite this repository to copy the citation in APA or BibTeX format.
A manuscript describing MethylSeg is in preparation. Its citation will be added when available.
Planned support
See ROADMAP.md
Reporting issues
To report a bug, request a feature, or ask a question about MethylSeg, open an issue on the GitHub issue tracker.
When reporting a bug, please include:
- Your MethylSeg version
- Your Python version and operating system
- The complete error message or traceback
- A minimal example that reproduces the problem, when possible
License
MethylSeg is distributed under the BSD 3-Clause License.
1 maintainer

Release files for methylseg 1.0.0
For a detailed explanation of source distributions (sdists) and built distributions (wheels), please see the package formats documentation.
Source distribution (sdist)
| File | Size | Uploaded | |
|---|---|---|---|
| methylseg-1.0.0.tar.gz | 76.1 kB | Details |
Built distribution (wheel)
| File | Interpreter | ABI | Platform | Reset |
|---|---|---|---|---|
| methylseg-1.0.0-py3-none-any.whl | Python 3 | none | any | Details |
Total release size: 155.1 kB
Release files / methylseg-1.0.0.tar.gz
| Download URL | methylseg-1.0.0.tar.gz |
|---|---|
| Size | 76.1 kB |
| Tags | Source |
|
SHA-256 checksum How to use checksums |
185cb15f3c4d681218e33e78c7d0cb2382beb96b5e71404a110a36c2c86cd44b
|
|
BLAKE2b-256 checksum How to use checksums |
7bf958a5130ce64dd13022b40ed18ff5faa27bd4e457717b8a15361c4ff1bdb2
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
Yes |
| Uploaded via |
twine/7.0.0 CPython/3.13.14
|
Provenance
Provenance describes where a file came from. On PyPI, provenance is shared via attestations, which provide a verifiable record of the build or publishing details. View details, limitations and caveats.


PyPI Publish Attestation
PyPI verified that this artifact, at this checksum, originated from the publisher listed below.
Signed by GitHub Actions, verified by PyPI on Sep 22, 2026.
Transparency logRelease files / methylseg-1.0.0-py3-none-any.whl
| Download URL | methylseg-1.0.0-py3-none-any.whl |
|---|---|
| Size | 78.9 kB |
| Tags | Python 3 |
|
SHA-256 checksum How to use checksums |
bdcd4e7f848145adf156671e3551714bade8a4ec86a8389b923060865862d912
|
|
BLAKE2b-256 checksum How to use checksums |
d0489ef6e240db5fcbd28d9aa57c35e0409f8e5156beec9ddf70896cf4d53d86
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
Yes |
| Uploaded via |
twine/7.0.0 CPython/3.13.14
|
Provenance
Provenance describes where a file came from. On PyPI, provenance is shared via attestations, which provide a verifiable record of the build or publishing details. View details, limitations and caveats.
PyPI Publish Attestation
PyPI verified that this artifact, at this checksum, originated from the publisher listed below.
Signed by GitHub Actions, verified by PyPI on Sep 22, 2026.
Transparency log













