

floria-strainer: strains out genomes from floria.
Introduction
Given the output of the strain haplotyping software floria1 , floria-strainer computes the allele frequency at each variable position of each haploset identified by floria to cluster them into the different strains composing the mixture, using a Gaussian Mixture Model.
Install
pip install floria-strainer
Quick start
Running floria-strainer on the provided test data
$ floria-strainer -b tests/data/test_short.bam tests/data/floria_out_dir
INFO - Writing the straining informations to floria_strained.strained.csv.
INFO - 2 strains were found: 0, 1
INFO - Writing the BAM file in tag mode to floria_strained.bam.
Visualizing the strain clustering method
-
In this example, short-reads are aligned to a reference genome (fig 1). At each variable position, we can observe a 2 different alleles, which in the case of this haploid organism, corresponds to a mixture of 2 different strains.
-
floria-strainer takes the output of floria, with reads having been assigned to a haploset (fig 2) based on the MEC criteria.
Expand to see a high level overview of floria
The haplotyping process floria is conceptually similar to the one of de novo assembly of reads into contigs, where instead of contigs, read haplosets are the results of the flow (ie. least costly path) optimization through the different reads represented as a graph. More details in the floria article 1.
- Based on the average allele frequency of each haploset, reads are clustered in the different strains. In this example (fig 3), there are two strains of minor and major allele frequency, which floria-strainer clustered in strain 0 and strain 1.
Reads that weren't assigned to any haploset by floria, or whose haploset do not cluster well enough are not assigned to any strain. They are considered to be shared by the different strains present in the alignment.
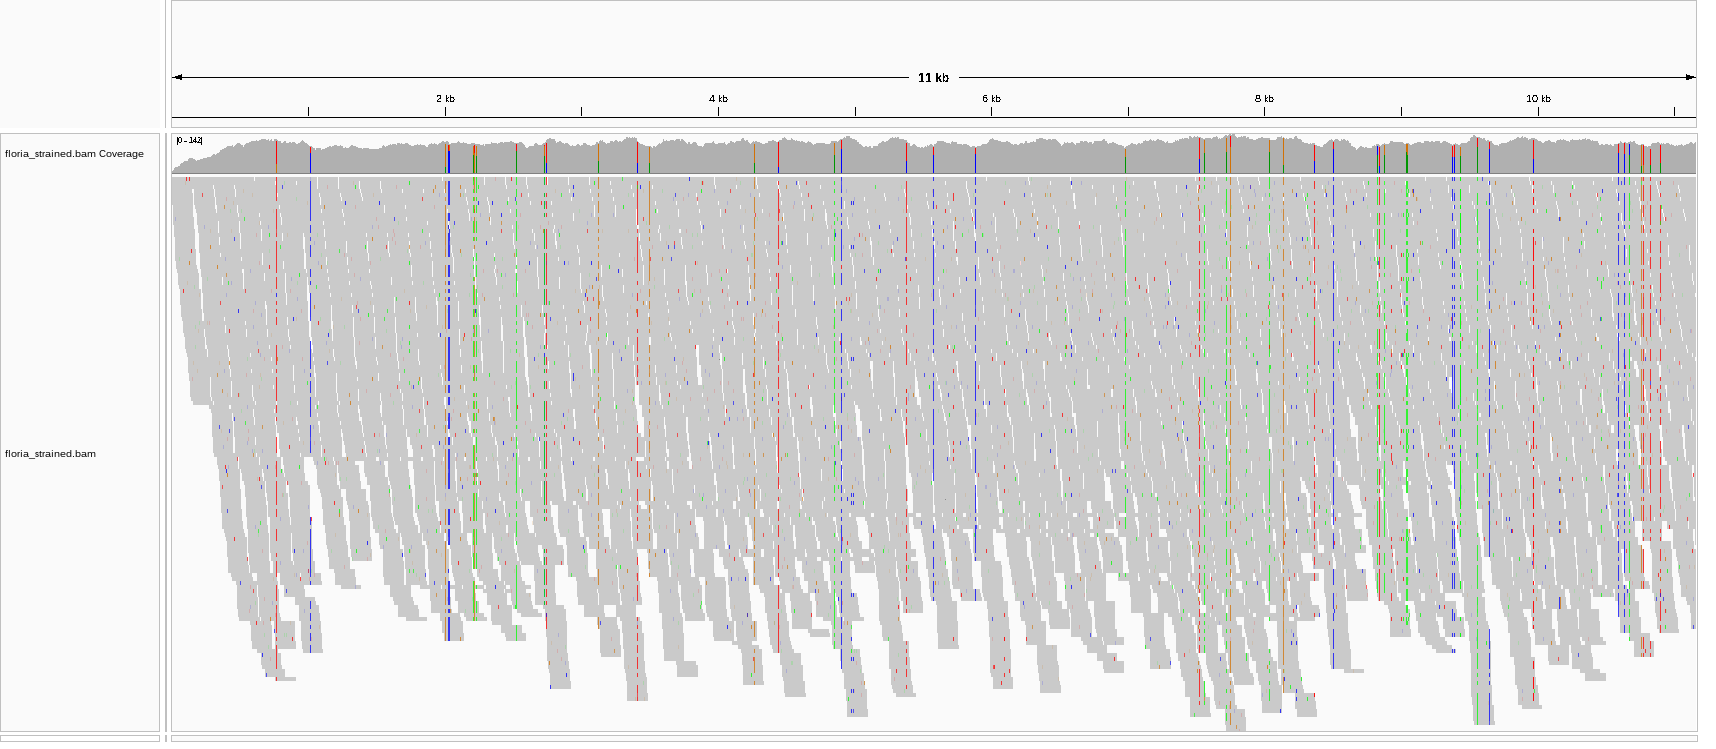
Fig 1: Reads aligned to the reference genome, visualized in IGV. The top track represents the reference genome, with variants indicated in the different colors.
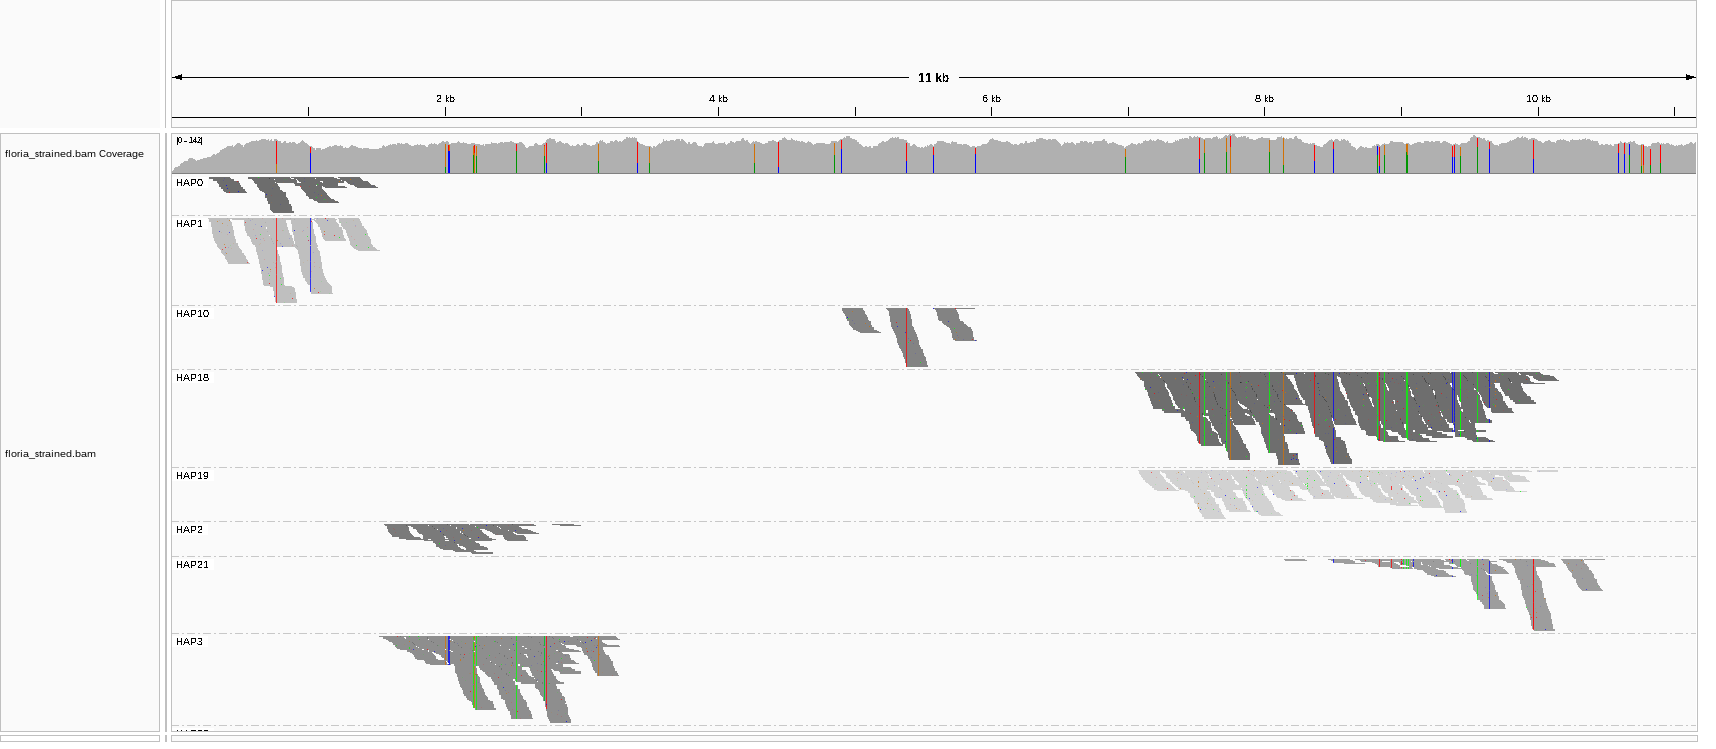
Fig 2: Reads are grouped and colored by the HP HaPloset tag as annotated by Floria.
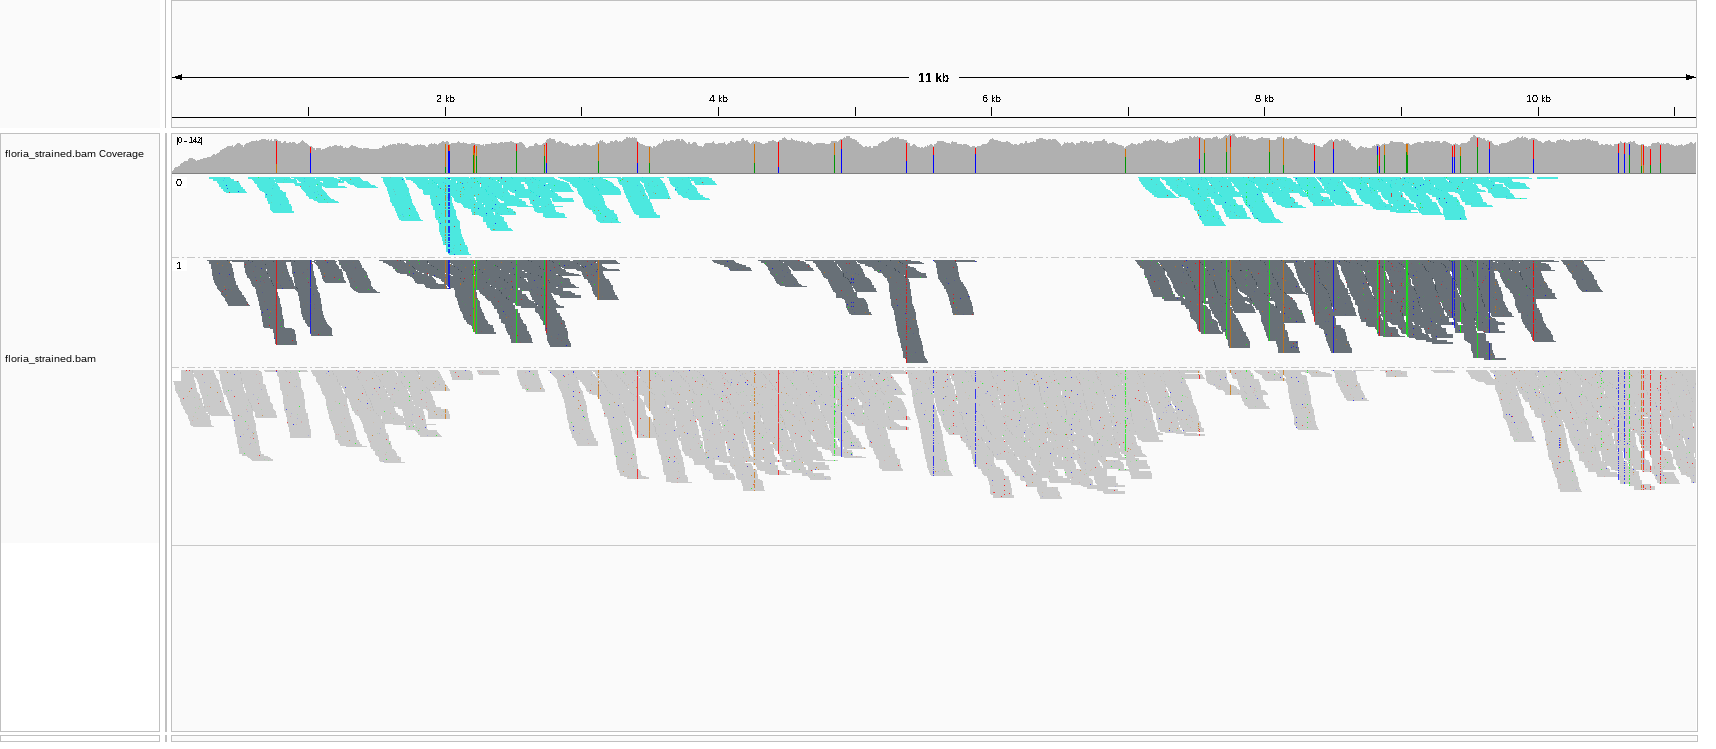
Fig 3: Reads are grouped and colored by the ST STrain tag as annotated by floria-strainer.
Help
$floria-strainer --help

 Usage: floria-strainer [OPTIONS] FLORIA_OUTDIR

 Strain the haplotypes in the floria output directory.
 Author: Maxime Borry

╭─ Options ────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────╮
│    --version                      Show the version and exit.                                                                                                             │
│    --nb-strains  -n  INTEGER      Number of strains to keep. If 0, the number of strains will be determined by the mean floria average strain count with HAPQ > 15.      │
│                                   [default: 0]                                                                                                                           │
│    --hapq-cut    -h  INTEGER      Minimum HAPQ threshold [default: 15]                                                                                                   │
│    --sp-cut      -s  FLOAT        Minimum strain clustering probability threshold [default: 0.5]                                                                         │
│ *  --bam         -b  PATH         Input BAM file [required]                                                                                                              │
│    --mode        -m  [tag|split]  BAM output mode. Tag: add ST (strain) tags to the reads. Split: split the reads in different BAM files per strain. [default: tag]      │
│ *  --basename    -o  TEXT         Output file basaneme [default: floria_strained] [required]                                                                             │
│    --help                         Show this message and exit.                                                                                                            │
╰──────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────────╯
Tests
After cloning/pulling repo
$ pip install poetry pytest
$ poetry run pytest -vv
1 maintainer

Metadata
1 maintainer
Release files for floria-strainer 0.2.1
For a detailed explanation of source distributions (sdists) and built distributions (wheels), please see the package formats documentation.
Source distribution (sdist)
| File | Size | Uploaded | Provenance | |
|---|---|---|---|---|
| floria_strainer-0.2.1.tar.gz | 19.9 kB | Without provenance | Details |
Built distribution (wheel)
| File | Interpreter | ABI | Platform | Reset |
|---|---|---|---|---|
| floria_strainer-0.2.1-py3-none-any.whl | Python 3 | none | any | Details |
Total release size: 40.3 kB
Release files / floria_strainer-0.2.1.tar.gz
| Download URL | floria_strainer-0.2.1.tar.gz |
|---|---|
| Size | 19.9 kB |
| Tags | Source |
|
SHA-256 checksum How to use checksums |
9680ee87186974c4cf2dc05f3bb0c3d5a2783e36673723bd9579da4658bacc91
|
|
BLAKE2b-256 checksum How to use checksums |
d88e19d71a7075d8e6ef63bf460de16b08772dcad1f2da68702511eb4d77cf88
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
No |
| Uploaded via |
poetry/1.8.3 CPython/3.12.1 Linux/6.5.0-1022-azure
|
Release files / floria_strainer-0.2.1-py3-none-any.whl
| Download URL | floria_strainer-0.2.1-py3-none-any.whl |
|---|---|
| Size | 20.4 kB |
| Tags | Python 3 |
|
SHA-256 checksum How to use checksums |
8239754c533b4bc70e1f6a21ab0f0384f67e859b9d9c9cbc035ed54dc30b0ea3
|
|
BLAKE2b-256 checksum How to use checksums |
019aaca56ae1320275a8efbf90645eb91f52592c9db526c7d2fb4a6343bc0347
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
No |
| Uploaded via |
poetry/1.8.3 CPython/3.12.1 Linux/6.5.0-1022-azure
|
Security for floria-strainer 0.2.1
Provenance
Provenance describes where a file came from. On PyPI, provenance is shared via attestations, which provide a verifiable record of the build or publishing details. View details, limitations and caveats.
No provenance found
None of the files in this release were uploaded with provenance metadata.
A note on package security
PyPI is a repository for distributing packages, not a service for auditing them. While third-party security researchers and our staff do their best to detect and remove malicious packages, PyPI cannot guarantee the quality, security or suitability of any given package.
To learn more about protecting yourself from malicious actors, visit our package security documentation.














