
Supply motif models and a DNA length. Motif Balance searches for sequences that strengthen the weakest desired match while optionally limiting unwanted matches. Inspect the recovered sites, choose different arrangements, then expand each layout into nucleotide alternatives.
Documentation · Supply motifs · Python API
Try a design
Fit the E. coli ArgR and Cra preferences into 25 bases. Their motif models span 25 and 14 positions, so the shorter match must fit within the same DNA as the wider one. These profiles were inferred from the DAP-seq data in Baumgart et al. (2021), Supplementary Data 2.
1. Install and prepare the profiles
With uv installed:
# Create a project and install the package from PyPI.
uv init --python 3.12 my-project
cd my-project
uv add 'motif-balance==0.8.1'

# Fetch the attributed example, its source record, and its editable design request.
MB_EXAMPLE_URL=https://raw.githubusercontent.com/e-south/motif-balance/v0.8.1/examples/argr-cra
curl -fLO "$MB_EXAMPLE_URL/prepare_inputs.py"
curl -fLO "$MB_EXAMPLE_URL/SOURCE.json"
curl -fLO "$MB_EXAMPLE_URL/design.yaml"

# Check the publisher's data and prepare local ArgR and Cra models.
uv run python prepare_inputs.py --out inputs
design.yaml requests four 25-base sequences, 4,096 candidate evaluations, and
seed 7. Edit that file to change the request. The
preparation record
explains the probability adjustment and source checks.
2. Design and inspect DNA
# Search for DNA with a strong weakest motif match, and save its sequences and scores.
uv run motif-balance design design.yaml --out result

# Draw the selected sites and logos, with the score and search summaries alongside.
uv run motif-balance inspect result --format html --out review.html
Open review.html. The best candidate scores about 0.855. The review shows
where each motif matches, on which strand, and how well it scores. Every candidate
is scanned on both strands.
3. Collect different arrangements
# Keep up to two representatives with different motif order, strand, or overlap.
uv run motif-balance collect result --count 2 --out collection.json
The representatives score about 0.855 and 0.801. Selection uses the retained search pool and reports any shortfall. The saved collection carries its sequences, motif models, and selected sites. Arrangement definitions explain which differences count.
4. Expand each arrangement into sequence alternatives
# Vary nucleotides within each layout while retaining its selected motif sites.
# Every returned sequence must have balance at least 0.8.
uv run motif-balance expand collection.json --all --min-balance 0.8 --out expanded
Each arrangement returns 256 sequences at the default output limit. Every
sequence meets the 0.8 floor and retains its parent's selected motif positions
and strands. Open the FASTA and per-motif score table for each arrangement in
expanded/; collection.json contains the complete record. Other designs can
yield fewer alternatives.
Sequence expansion explains the score floor and work limits. For synthesis as a checked ambiguity template, use the separate degenerate-library operation.
Use a new output name when repeating a step. The Python tutorial uses the same example in code. Installation covers pip and optional video export.
Inspect a larger design
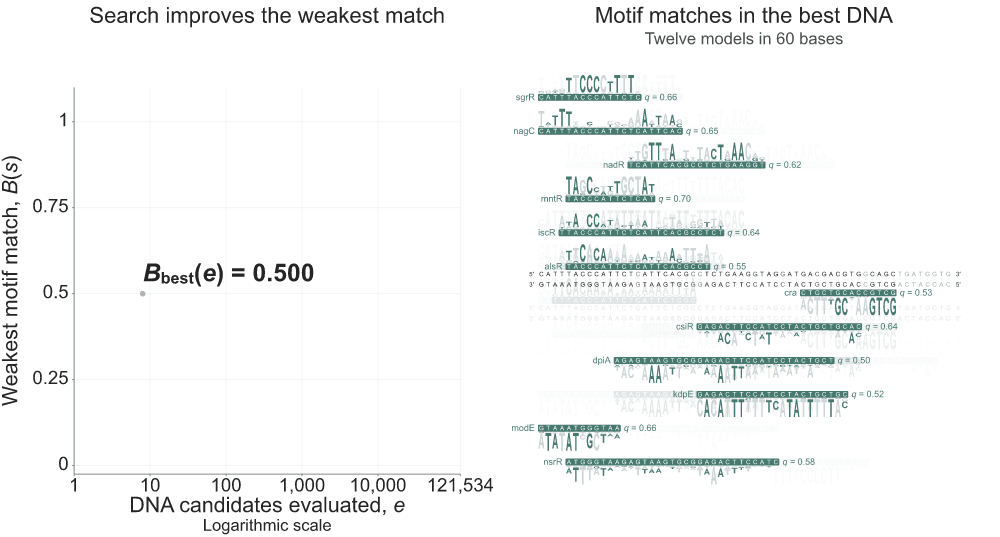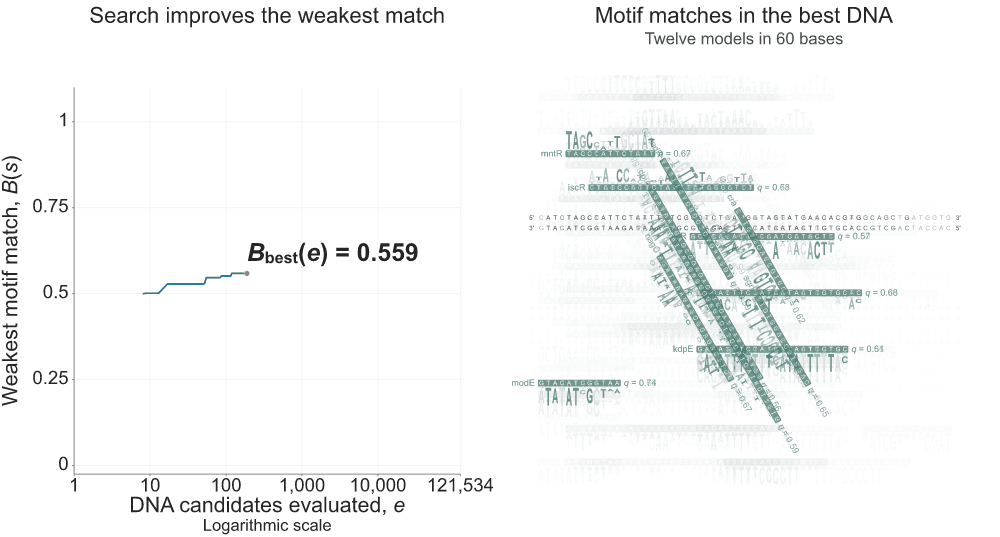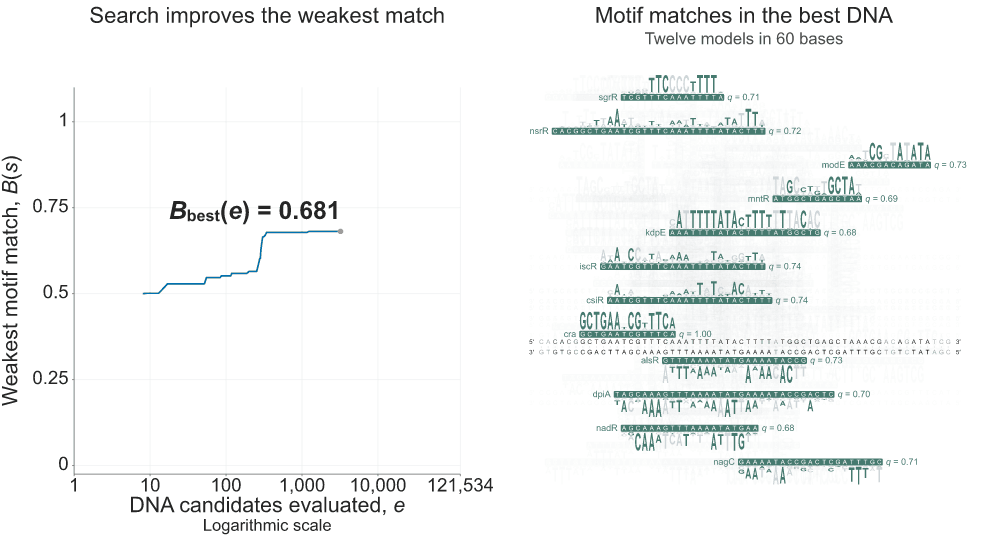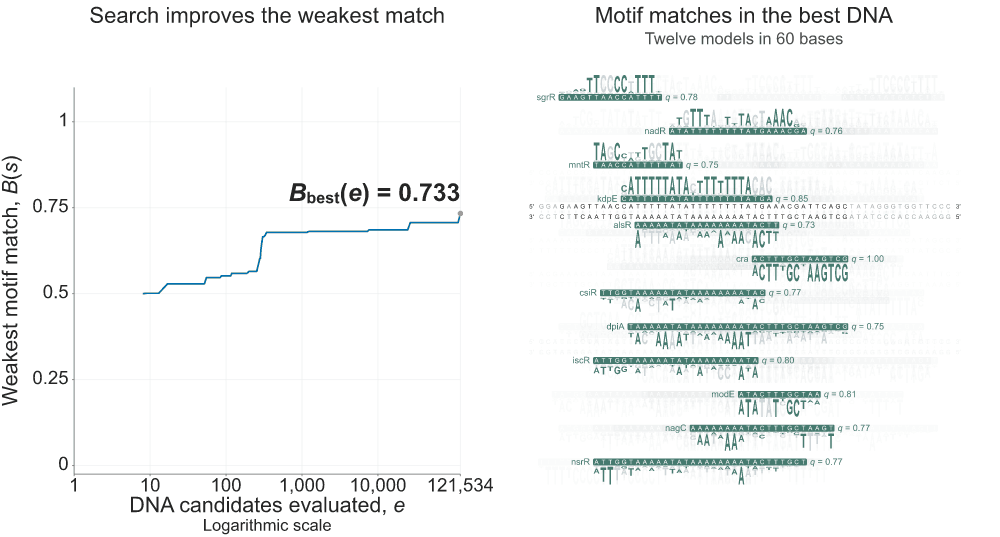
The twelve-model example shows improvements during a recorded search in 60-base DNA. The chart follows the best balance found. Colored motif windows show that sequence; faint gray windows show recorded search candidates. Transitions connect saved placements.
Python 3.12–3.14 · Linux and macOS · MIT license · Contributing
For study data, reproduction, and citation, use the Motif Balance Study. Publication DOI: pending.
1 maintainer

Metadata
1 maintainer
Release files for motif-balance 0.8.1
For a detailed explanation of source distributions (sdists) and built distributions (wheels), please see the package formats documentation.
Source distribution (sdist)
| File | Size | Uploaded | |
|---|---|---|---|
| motif_balance-0.8.1.tar.gz | 22.8 MB | Details |
Built distribution (wheel)
| File | Interpreter | ABI | Platform | Reset |
|---|---|---|---|---|
| motif_balance-0.8.1-py3-none-any.whl | Python 3 | none | any | Details |
Total release size: 23.0 MB
Release files / motif_balance-0.8.1.tar.gz
| Download URL | motif_balance-0.8.1.tar.gz |
|---|---|
| Size | 22.8 MB |
| Tags | Source |
|
SHA-256 checksum How to use checksums |
17ec7ea300af3f08610509e110545f2c391724a9ba8b237cfaf6169e754e7579
|
|
BLAKE2b-256 checksum How to use checksums |
6c6a733e904d5cb7cf5e885d026ab418f59af8e7a76522bcf9ad05edeb73b492
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
Yes |
| Uploaded via |
twine/7.0.0 CPython/3.13.14
|
Provenance
Provenance describes where a file came from. On PyPI, provenance is shared via attestations, which provide a verifiable record of the build or publishing details. View details, limitations and caveats.


PyPI Publish Attestation
PyPI verified that this artifact, at this checksum, originated from the publisher listed below.
Signed by GitHub Actions, verified by PyPI on Oct 1, 2026.
Transparency logRelease files / motif_balance-0.8.1-py3-none-any.whl
| Download URL | motif_balance-0.8.1-py3-none-any.whl |
|---|---|
| Size | 211.1 kB |
| Tags | Python 3 |
|
SHA-256 checksum How to use checksums |
57fd6d6877ac4908e69ccacffd192e808f6d6c077d3eacbbf9ef332008f71ed0
|
|
BLAKE2b-256 checksum How to use checksums |
dea3f89af8a1680e428df6ac40fa7ca1344f247939dff458ff7ca640d159686d
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
Yes |
| Uploaded via |
twine/7.0.0 CPython/3.13.14
|
Provenance
Provenance describes where a file came from. On PyPI, provenance is shared via attestations, which provide a verifiable record of the build or publishing details. View details, limitations and caveats.
PyPI Publish Attestation
PyPI verified that this artifact, at this checksum, originated from the publisher listed below.
Signed by GitHub Actions, verified by PyPI on Oct 1, 2026.
Transparency log













