


Quasigraph is an open-source toolkit designed for generating chemical and geometric descriptors to be used in machine learning models.
Installation
The easiest method to install quasigraph is by utilizing pip (Python 3.10 or newer):
$ pip install quasigraph
Optional extras: quasigraph[ml] (scikit-learn transformer), quasigraph[pymatgen] (pymatgen input), quasigraph[graph] (networkx export), quasigraph[mendeleev] (get_element), quasigraph[dev] (tests, ruff, mypy).
Getting started
from ase.build import molecule
from quasigraph import QuasiGraph

# Initialize an Atoms object for methanol (CH3OH) using ASE's molecule function
atoms = molecule('CH3OH')

# Instantiate a QuasiGraph object containing chemical and coordination numbers
qgr = QuasiGraph(atoms)

# Convert the QuasiGraph object into a pandas DataFrame
df = qgr.get_dataframe()

# Convert the QuasiGraph object into a vector
vector = qgr.get_vector()

# Descriptor of the whole structure, independent of the order and number of atoms
features = qgr.get_structure_features()
Structures written by DFT codes (VASP, GPAW, Quantum ESPRESSO, SIESTA, ...) are read with ase.io.read; pymatgen Structure and Molecule objects are accepted as well.
Descriptor
The descriptor can be separated into two parts, a chemical part and a geometric part.
Chemical part
The chemical part of the descriptor uses element properties from the Mendeleev library, bundled with quasigraph as a table. By default it holds the valence electron concentration (VEC), atomic radius (Å), Pauling electronegativity and electron affinity (eV) of every atom; chemical_features selects others among AVAILABLE_FEATURES (group, period, atomic_weight, covalent_radius, atomic_radius, vdw_radius, en_pauling, electron_affinity, dipole_polarizability, VEC).
For example, for methanol (CH3OH) we have the table:
| VEC | atomic_radius | en_pauling | electron_affinity | |
|---|---|---|---|---|
| 0 | 4 | 0.7 | 2.55 | 1.26212 |
| 1 | 6 | 0.6 | 3.44 | 1.46111 |
| 2 | 1 | 0.25 | 2.2 | 0.754195 |
| 3 | 1 | 0.25 | 2.2 | 0.754195 |
| 4 | 1 | 0.25 | 2.2 | 0.754195 |
| 5 | 1 | 0.25 | 2.2 | 0.754195 |
Geometric part
The geometric part involves identifying all bonds and computing the coordination numbers for each atom, indicated as CN. Additionally, the generalized coordination number (GCN)1 is determined by summing the coordination numbers of the neighboring ligands for each atom and normalizing this sum by the highest coordination number found in the molecule. To compare GCN values between structures, fix the normalization instead with cn_max, e.g. QuasiGraph(atoms, cn_max=12) for fcc metals as in the original definition.
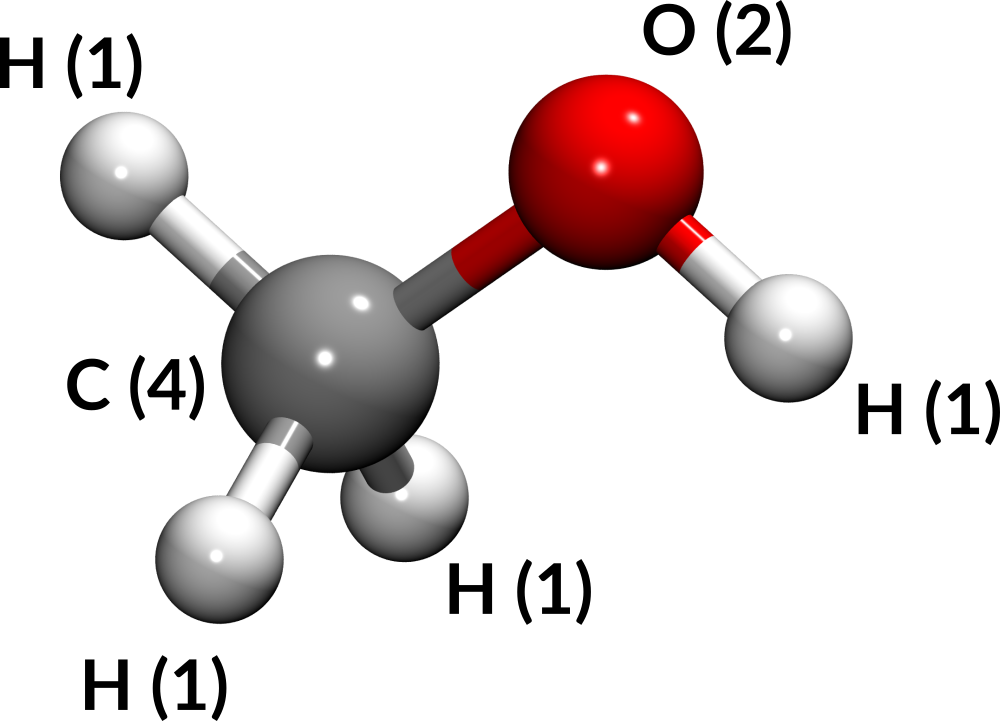
Figure 1 - Schematic representation of the methanol molecule, indicating the chemical symbol and coordination number (CN) for every atom.
For example, for methanol (CH3OH) we have the geometric data, as shown in Fig. 1.
| CN | GCN |
|---|---|
| 4 | 1.25 |
| 2 | 1.25 |
| 1 | 1.00 |
| 1 | 0.50 |
| 1 | 1.00 |
| 1 | 1.00 |
Periodic structures
Periodic boundary conditions are taken from atoms.pbc unless the pbc argument is given (e.g. pbc=False treats a periodic cell as an isolated cluster). Along each periodic axis, as many periodic images are included as needed to find every bond, also for small or strongly skewed cells. Periodic images of a neighbour count separately in CN, GCN and the neighbour shells of get_site_environment, so a primitive cell and any supercell of it give the same descriptor.
Chemical properties missing from the Mendeleev database (e.g. the Pauling electronegativity of noble gases) are returned as NaN, with a MissingFeatureWarning.
Large structures: backend="vesin"
By default, bonds are found from the full distance matrix (a tensor over periodic images for a periodic structure), whose memory grows with the square of the number of atoms. With backend="vesin" they are found with the sparse neighbour list of the vesin library instead, which scales linearly:
qgr = QuasiGraph(atoms, backend="vesin")
Both backends give the same descriptor (CN_smooth agrees to ~1e-8, since the vesin sum of its sigmoid stops 20 temperature beyond the bond threshold). With "vesin" the dense attributes distances, distances_tensor, bonds and bond_threshold are not built; the bonds are in pairs instead. For a periodic 2916-atom Pt supercell with all geometric features, "dense" takes 24 s and 9.5 GB, "vesin" 0.07 s; for a ~55-atom nanoparticle both take a few milliseconds ("dense" is slightly faster).
Continuous bond information (optional)
By default the geometric part is CN and GCN. The geometric_features argument selects any of the per-atom features below; the chemical columns come first, then the geometric ones in the order given.
from quasigraph import QuasiGraph, GEOMETRIC_FEATURES

qgr = QuasiGraph(atoms, geometric_features=GEOMETRIC_FEATURES)
qgr.get_dataframe()   # ... CN, GCN, CN_smooth, bond_mean, bond_min, bond_max, bond_std, bond_strain
| Feature | Meaning |
|---|---|
CN |
number of bonded atoms (a pair is bonded when its distance is at most (1 + tolerance) times the sum of covalent radii) |
GCN |
generalized coordination number |
CN_smooth |
continuous CN: every pair contributes a sigmoid weight 1 / (1 + exp((d − rc) / T)) centred on the bond threshold rc = (1 + tolerance)(ri + rj), with temperature T in Å (default 0.1). T → 0 gives a step function and CN_smooth = CN |
bond_mean, bond_min, bond_max, bond_std |
statistics of the atom's bond lengths (0 for an atom without bonds) |
bond_strain |
mean of (bond length / sum of covalent radii) − 1: positive for stretched bonds |
Covalent radii are the Pyykkö values from Mendeleev. Metallic bonds are typically 10–20 % longer than the sum of these radii, so in a perfect fcc metal bond_strain ≈ 0.15; the useful information is the variation between sites. With the default temperature of 0.1 Å a bond 0.3 Å inside the threshold has weight 0.95 and one 0.3 Å outside it 0.05, so CN_smooth stays close to CN while varying continuously with bond length; a larger temperature smooths more.
Adsorption-site environment
For a target that depends on one atom, such as the adsorption free energy of a hydrogen atom, get_site_environment aggregates the descriptor over the neighbour shells of that atom on the bond graph and returns a fixed-length, named vector that does not depend on the number of atoms or their order:
qgr = QuasiGraph(atoms, geometric_features=GEOMETRIC_FEATURES)
env = qgr.get_site_environment("H", shells=2, elements=["Ag", "Au", "Cu", "Pd", "Pt"])  # pandas Series
vector = qgr.get_site_vector("H", shells=2, elements=["Ag", "Au", "Cu", "Pd", "Pt"])   # numpy array
site is an atom index or a chemical symbol that occurs once in the structure. The Series contains site_<feature> for the site atom itself and, for each shell k (atoms k bonds away from the site): shell<k>_n, shell<k>_n_<El> (counts per element in elements), shell<k>_dist_mean/min/max (distance from the site), shell<k>_<feature>_mean for chemical features and shell<k>_<feature>_mean/min/max for geometric features. Pass the same elements list for every structure of a dataset so all vectors have the same length; an empty shell contributes zeros.
Whole-structure descriptor
get_structure_features returns a named pandas Series that depends neither on the order nor on the number of atoms: n_atoms, the composition fractions frac_<El> and the mean, min, max and std of every per-atom feature (get_structure_vector gives the same as a numpy array). Pass the same elements list for every structure of a dataset.
qgr = QuasiGraph(atoms, geometric_features=GEOMETRIC_FEATURES)
qgr.get_structure_features(elements=["Ag", "Au", "Cu", "Pd", "Pt"])   # n_atoms, frac_Ag, ..., GCN_mean, GCN_min, ...
The flattened per-atom vector get_vector() depends on the order of the atoms and, unless nmax pads it, on their number; get_feature_names() names its entries.
Machine learning workflows
featurize computes a descriptor for every structure of a dataset and returns one DataFrame, with the same columns for every row (elements defaults to all elements of the dataset):
from ase.io import read
from quasigraph import featurize, GEOMETRIC_FEATURES

structures = [read(f) for f in files]
X = featurize(structures, kind="site", site="H", geometric_features=GEOMETRIC_FEATURES)   # or kind="structure" / "atoms"
n_jobs spreads the work over processes; it pays off for large structures, while for small ones (e.g. ~55-atom nanoparticles) starting the processes costs more than it saves. On macOS and Windows, call it under if __name__ == "__main__":.
QuasiGraphFeaturizer (pip install quasigraph[ml]) is a scikit-learn transformer, so quasigraph can be the first step of a pipeline and be cross-validated or tuned with the model:
from sklearn.ensemble import RandomForestRegressor
from sklearn.model_selection import cross_val_score
from sklearn.pipeline import make_pipeline
from quasigraph import QuasiGraphFeaturizer

model = make_pipeline(QuasiGraphFeaturizer(kind="site", site="H", cn_max=12), RandomForestRegressor())
scores = cross_val_score(model, structures, y, cv=5)
fit fixes the element list from the training structures; get_feature_names_out() names the columns and set_output(transform="pandas") returns DataFrames.
Bond graph
get_bonds() returns the bonds as an edge list (i, j, lattice translation shift_a/b/c, distance; every bond in both directions), and to_networkx() the bond graph as a networkx.MultiGraph with the per-atom features as node attributes (pip install quasigraph[graph]).
Package structure
quasigraph.quasigraph.QuasiGraph– user-facing class; holds an ASEAtomsobject asatoms(it is no longer anAtomssubclass) and exposesget_dataframe()/get_vector(),get_structure_features(),get_site_environment(),get_bonds()plus the attributescn,gcn,bonded_atoms,adjacencyand, with the dense backend,bonds,distances(non-periodic) anddistances_tensor(periodic).quasigraph.geometry– pure NumPy functions for distances, bonds, CN and GCN (distance_matrix,distance_tensor,bonds_from_distance_matrix,coordination_numbers,generalized_coordination_numbers, ...). They take plain arrays and can be reused without ASE.quasigraph.neighbors– the same features computed from a sparse pair list found with vesin (neighbor_pairs,coordination_numbers,bond_length_statistics, ...), used bybackend="vesin".quasigraph.elements– chemical features (get_feature) from the bundled tablequasigraph/data/elements.json, built from the Mendeleev library byscripts/build_element_table.py, so results do not depend on the installed Mendeleev version.quasigraph.ptable– hand-curated tables (VEC,CONFIG) not available in Mendeleev.quasigraph.batch–featurize, descriptors of a dataset.quasigraph.estimators–QuasiGraphFeaturizer, the scikit-learn transformer.
Development
$ pip install -e ".[dev]"
$ pytest          # tests
$ ruff check .    # lint
$ mypy            # type check
Changes are listed in CHANGELOG.md.
License
This is an open source code under MIT License.
Acknowledgements
We thank financial support from FAPESP (Grant No. 2022/14549-3), INCT Materials Informatics (Grant No. 406447/2022-5), and CNPq (Grant No. 311324/2020-7).
-
Calle-Vallejo, F., Martínez, J. I., García-Lastra, J. M., Sautet, P. & Loffreda, D. Fast Prediction of Adsorption Properties for Platinum Nanocatalysts with Generalized Coordination Numbers, Angew. Chem. Int. Ed. 53, 8316-8319 (2014). ↩
1 maintainer

Metadata
1 maintainer
Release files for quasigraph 26.10.1
For a detailed explanation of source distributions (sdists) and built distributions (wheels), please see the package formats documentation.
Source distribution (sdist)
| File | Size | Uploaded | |
|---|---|---|---|
| quasigraph-26.10.1.tar.gz | 191.3 kB | Details |
Built distribution (wheel)
| File | Interpreter | ABI | Platform | Reset |
|---|---|---|---|---|
| quasigraph-26.10.1-py3-none-any.whl | Python 3 | none | any | Details |
Total release size: 225.1 kB
Release files / quasigraph-26.10.1.tar.gz
| Download URL | quasigraph-26.10.1.tar.gz |
|---|---|
| Size | 191.3 kB |
| Tags | Source |
|
SHA-256 checksum How to use checksums |
78fc693a5267eca434f3885b97866f0c3eb4314b3b4a1dce975559cec1b567a1
|
|
BLAKE2b-256 checksum How to use checksums |
4bd59b45961825a06dc0d08410f109279990323292dde01023054179ba9694e1
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
No |
| Uploaded via |
twine/7.0.0 CPython/3.12.7
|
Release files / quasigraph-26.10.1-py3-none-any.whl
| Download URL | quasigraph-26.10.1-py3-none-any.whl |
|---|---|
| Size | 33.8 kB |
| Tags | Python 3 |
|
SHA-256 checksum How to use checksums |
664032c274d54e23acbe23760b6e1745a05e6d0714ed82edd4a31190f275fdae
|
|
BLAKE2b-256 checksum How to use checksums |
1b6c9701387fa810d14bc6f4522faa29ac33674ab74bb434ebe03960736622c4
|
| Upload date | |
|
Uploaded using Trusted Publishing? What is trusted publishing? |
No |
| Uploaded via |
twine/7.0.0 CPython/3.12.7
|














